Enantioselective catalytic desymmetric ester aminolysis
Wen-Ya Zheng
1,#
,
Miao He
1,#
,
Hua Wu
1,*

,
Yu-Ping He
2,*
*Correspondence to:
Hua Wu, Shanghai Key Laboratory for Molecular Engineering of Chiral Drugs, School of Pharmaceutical Sciences, Shanghai Jiao Tong University, Shanghai 200240, China.
E-mail: hua.wu@sjtu.edu.cn
Yu-Ping He, Department of Chemistry, College of Sciences, Shanghai University, Shanghai 200444, China. E-mail: yuping_he@shu.edu.cn
Yu-Ping He, Department of Chemistry, College of Sciences, Shanghai University, Shanghai 200444, China. E-mail: yuping_he@shu.edu.cn
Chiral Chem. 2027;3:202628. 10.70401/cc.2026.0037
Received: June 13, 2026Accepted: August 10, 2026Published: August 10, 2026
Abstract
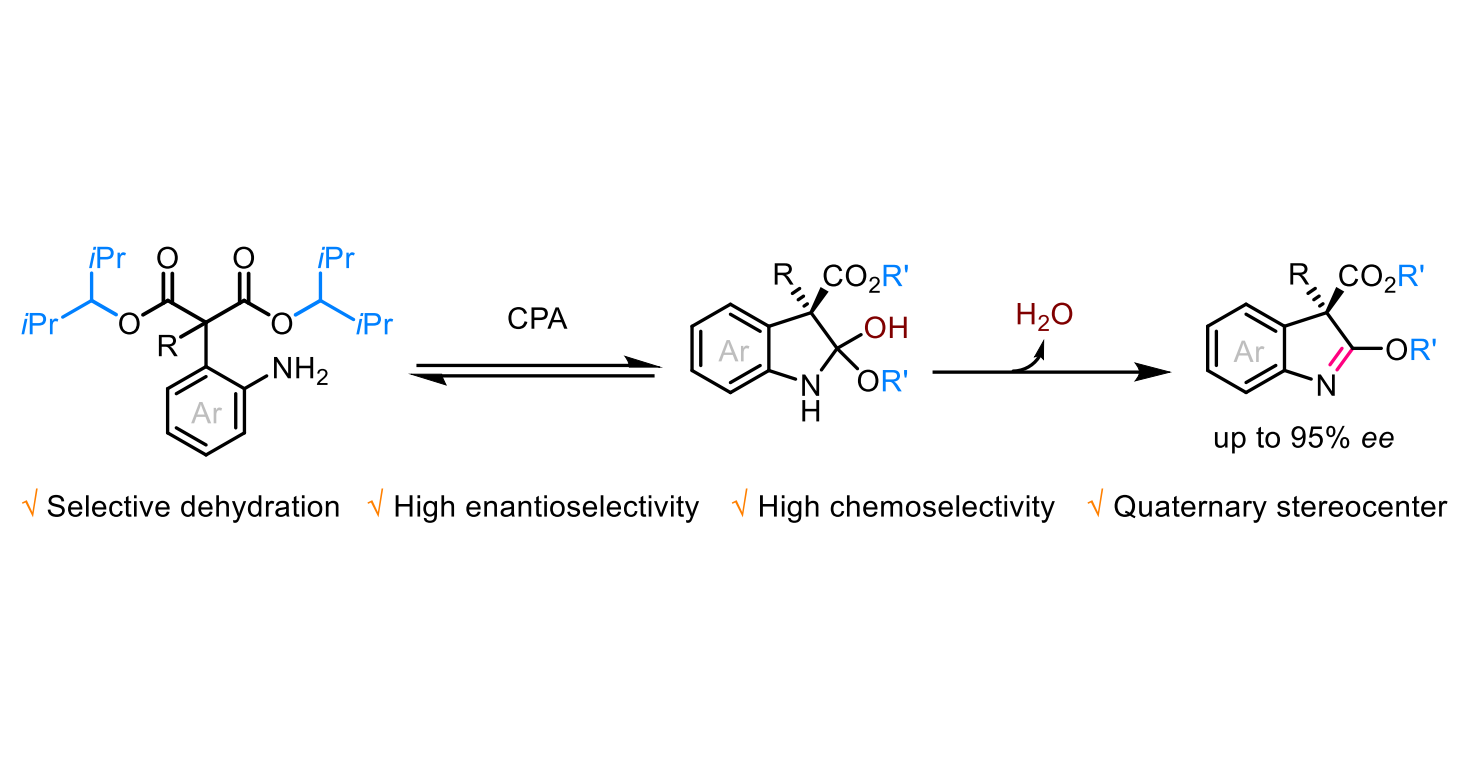
We report an organocatalytic enantioselective desymmetrization of 2-aminophenyl malonates enabled by intramolecular ester aminolysis. This protocol efficiently delivers C3-disubstituted oxindole derivatives bearing quaternary stereocenters under mild conditions with excellent chemo- and enantioselectivities. Notably, this reaction follows an acid-promoted dehydration pathway instead of an alcohol elimination pathway, which differs from conventional ester aminolysis reactions reported previously. Gram-scale synthesis and further derivatization of the resulting products demonstrate the synthetic utility of this transformation.
Graphical Abstract
Keywords
Desymmetrization, organocatalysis, quaternary stereocenter, ester aminolysis, oxindole
1. Introduction
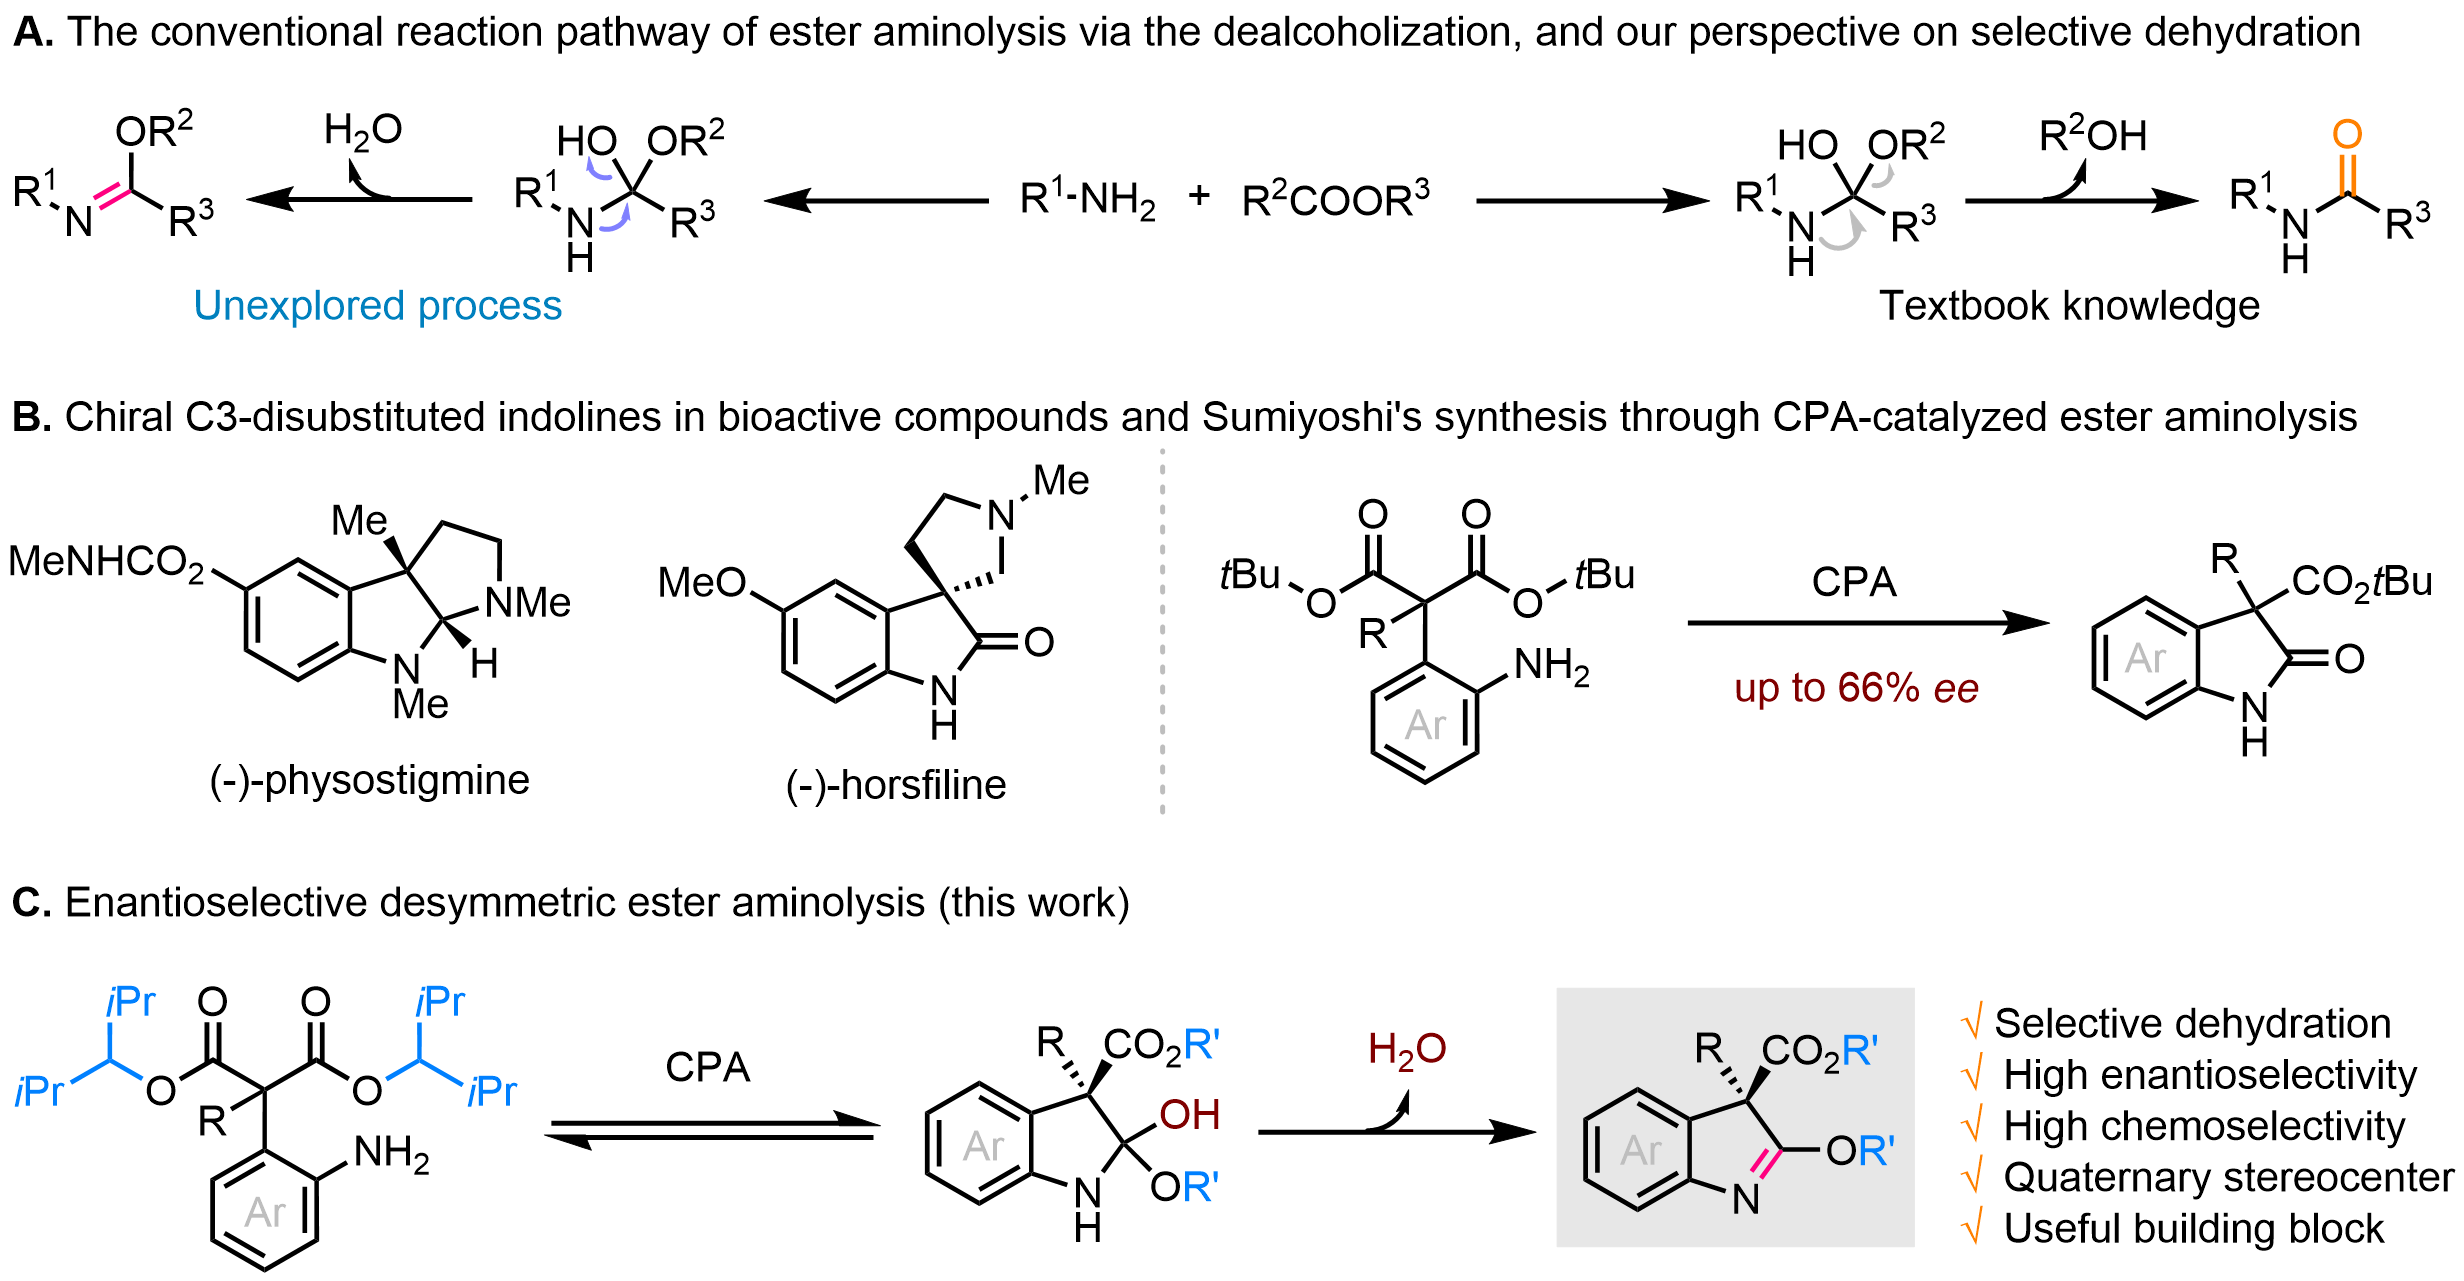
Ester aminolysis is one of the most fundamental and widely practiced reactions in organic synthesis, serving as the cornerstone for the preparation of amide bonds in pharmaceuticals, agrochemicals, and functional materials[1]. Conventionally, this transformation proceeds exclusively via the textbook-established dealcoholization pathway: the amine nucleophile first attacks the carbonyl group of the ester to generate a tetrahedral intermediate[2]. This intermediate then undergoes elimination of alcohol to afford the final amide product, a selectivity pattern so dominant that it is rarely questioned in standard synthetic design. By contrast, an alternative dehydration pathway from the identical tetrahedral intermediate remains completely unexplored. In this pathway, the intermediate would lose water instead of alcohol to form an imine product, a transformation that would enable direct access to valuable imine scaffolds from readily available ester and amine feedstocks, bypassing the need for pre-synthesized carbonyl substrates (Scheme 1A). Despite its synthetic appeal, this elusive selectivity switch has never been realized, likely due to the strong thermodynamic preference for alcohol elimination in the absence of specific reaction control. Furthermore, its acid-catalyzed asymmetric variant, which would provide a straightforward route to enantioenriched chiral imines, has not been reported to date.
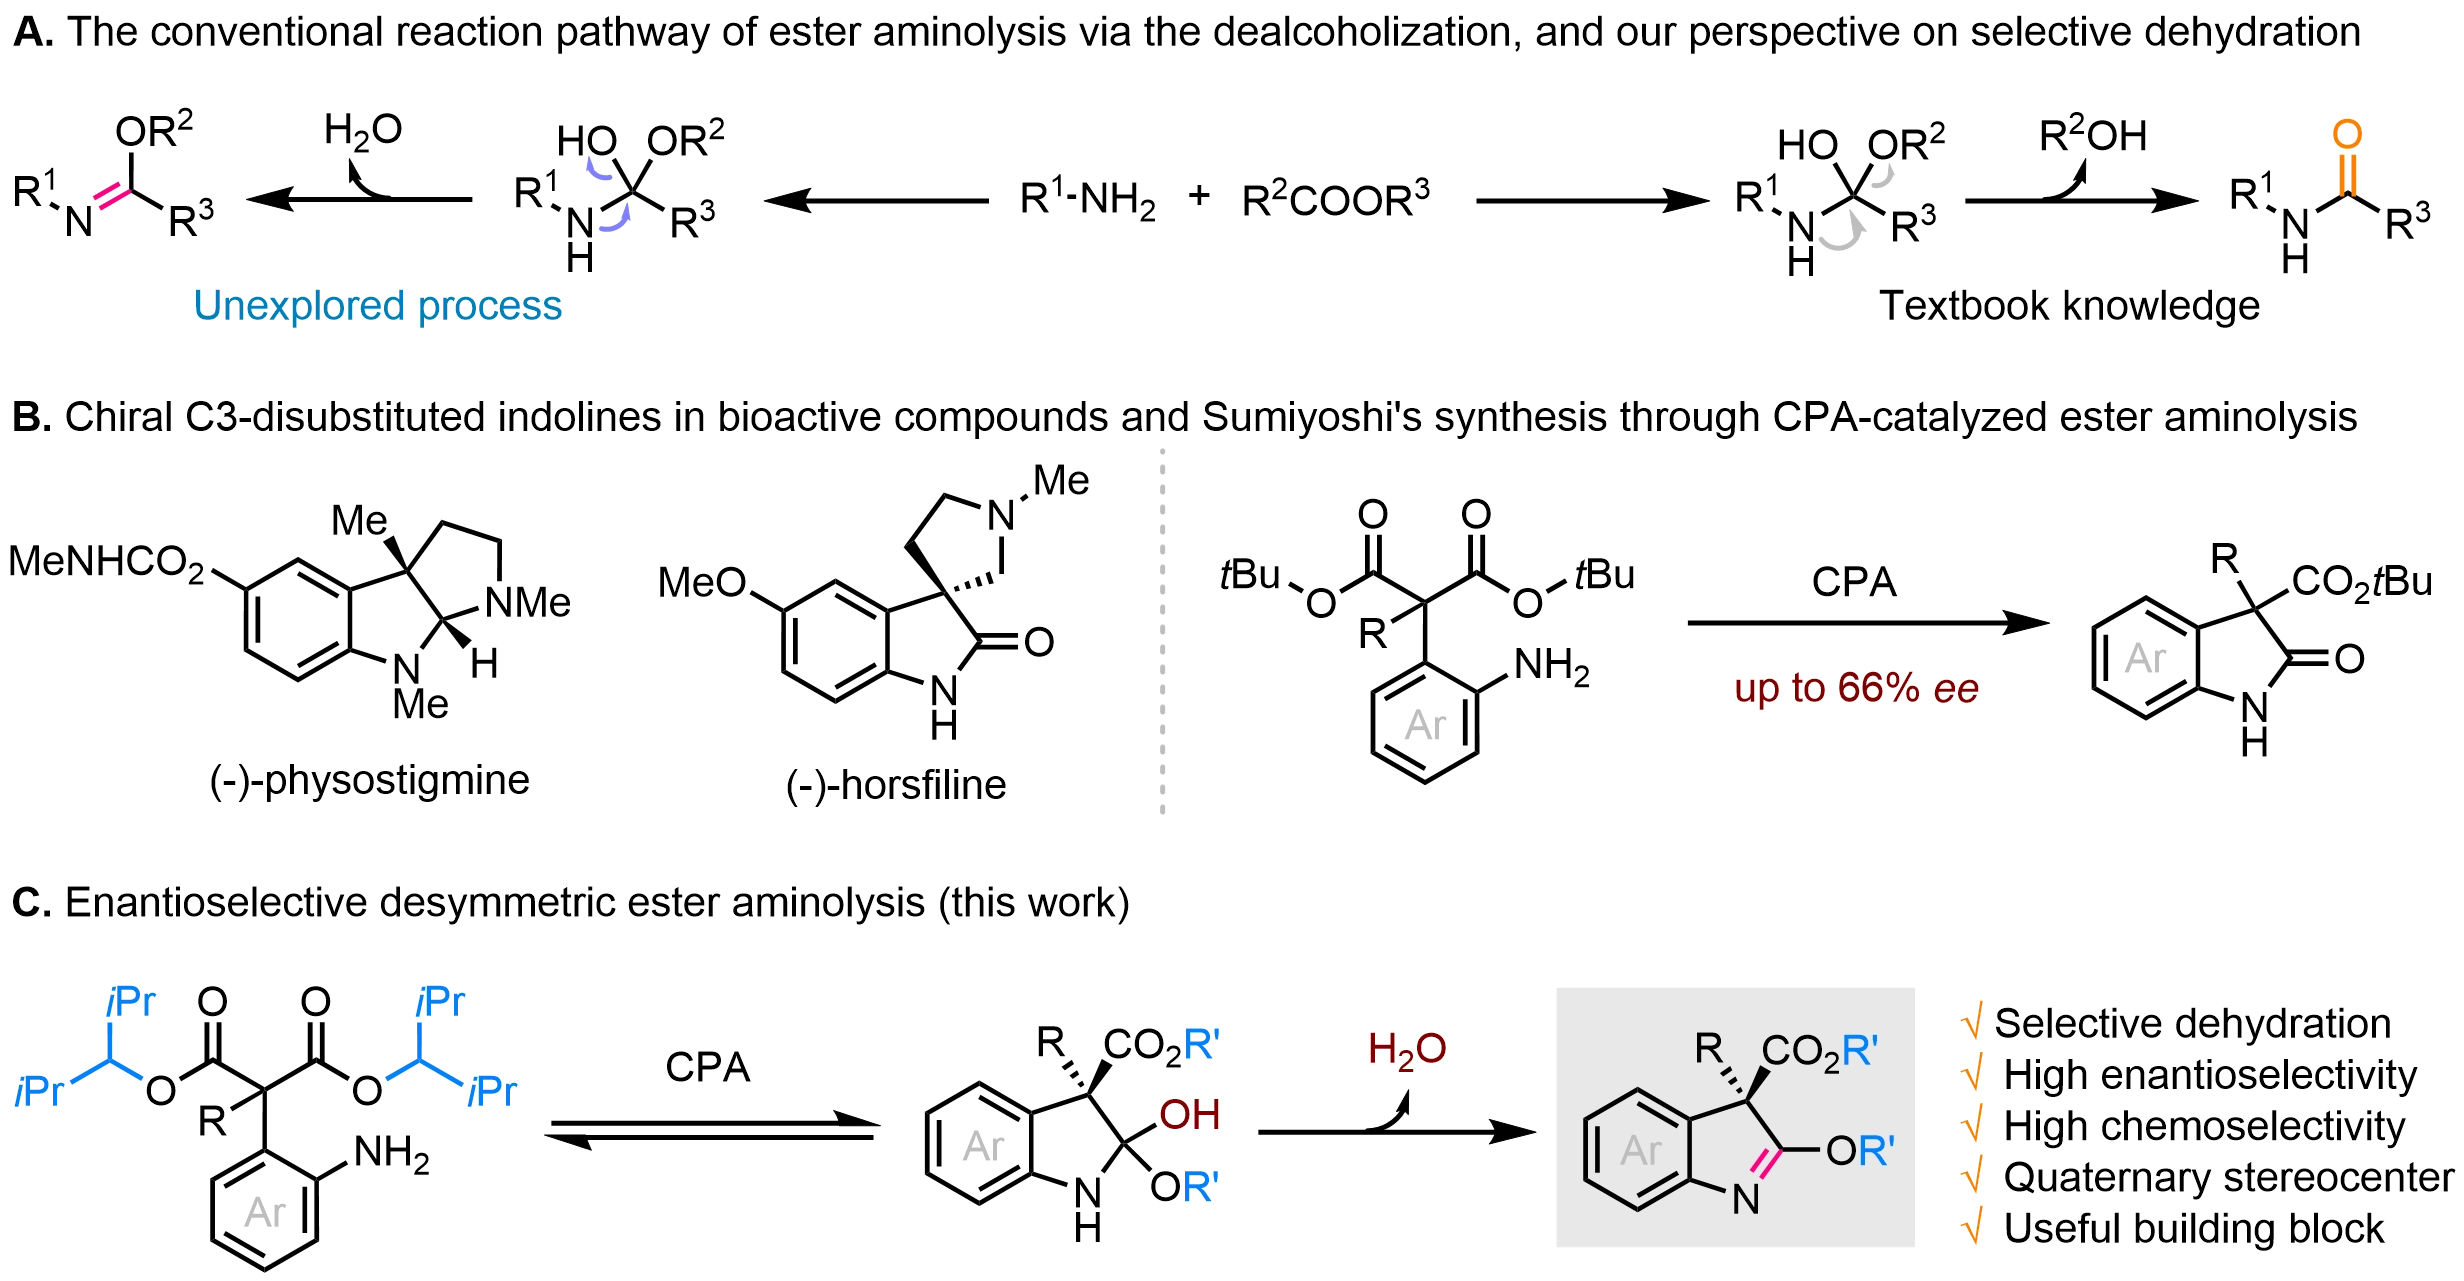
Chiral C3-disubstituted oxindole skeletons are ubiquitous core structures in numerous bioactive natural alkaloids and pharmaceutical molecules[3]. On the other hand, a desymmetrization strategy is a powerful synthetic tool for the construction of all-carbon quaternary stereocenters[4,5]. However, asymmetric catalytic synthesis of C3-disubstituted oxindole derivatives through a desymmetrization approach has rarely been reported, especially by organocatalysis. In this regard, Liu, Kwon and coworkers reported a transition-metal catalyzed desymmetrizing Staudinger–aza–Wittig reaction that enabled enantioselective construction of 3,3-disubstituted indolin-2-ones[6]. Sumiyoshi and co-workers reported a desymmetrizing lactamization of di-tert-butyl 2-alkyl-2-(2-aminophenyl)malonates via intramolecular ester aminolysis catalyzed by chiral phosphoric acid[7]. This transformation delivered C3-substituted oxindoles, yet extensive screening afforded only a maximum enantiomeric excess of 66% (Scheme 1B).
In connection with our ongoing research interest in the enantioselective desymmetrization of malonate derivatives[8], we herein present an organocatalytic ester aminolysis reaction delivering C3-quaternary oxindole derivatives (Scheme 1C). This transformation preferentially follows a dehydration pathway instead of the conventional dealcoholation pathway, and exhibits excellent enantioselectivity, high chemoselectivity, as well as a good substrate scope.
2. Experimental section
Compounds 3. A suspension of 1 (0.05 mmol, 1.0 equiv.), 2h (10 mol%), and 5 Å molecular sieves (15.0 mg) in CHCl3 (0.5 mL) was stirred in a dry sealed tube under argon. The reaction mixture was stirred at 25 °C for the indicated time under argon. After completion of the reaction (monitored by TLC), the solvent was removed in vacuo. The residue was purified by column chromatography on silica gel, eluting with petroleum ether/ethyl acetate (10:1, v/v) to afford the products 3 in 80-98% yields with 61-95% ee.
Compound 4b. 6 N HCl (0.075 mmol, 1.5 equiv.) was added to the solution of 3b (0.05 mmol, 1.0 equiv.) in THF (0.5 mL) at room temperature. The mixture was stirred for 3 h at room temperature. After completion of the reaction (monitored by TLC), the solvent was removed in vacuo. The residue was purified by column chromatography on silica gel, eluting with petroleum ether/ethyl acetate (10:1, v/v) to give compound 4b in 94% yield (19.5 mg) with 89% ee as a white solid.
Compound 5. DIBAL-H (1.0 M in toluene, 0.1 mmol, 2.0 equiv.) was added to a solution of 4b (0.05 mmol, 1.0 equiv.) in anhydrous DCM (0.5 mL) at -40 °C. The mixture was stirred at the same temperature for 2 h. After completion of the reaction (monitored by TLC), the solvent was removed in vacuo. The residue was purified by column chromatography on silica gel, eluting with ethyl acetate to give compound 5 in 70% yield (7.2 mg) with 87% ee as a colorless oil.
3. Results and Discussion
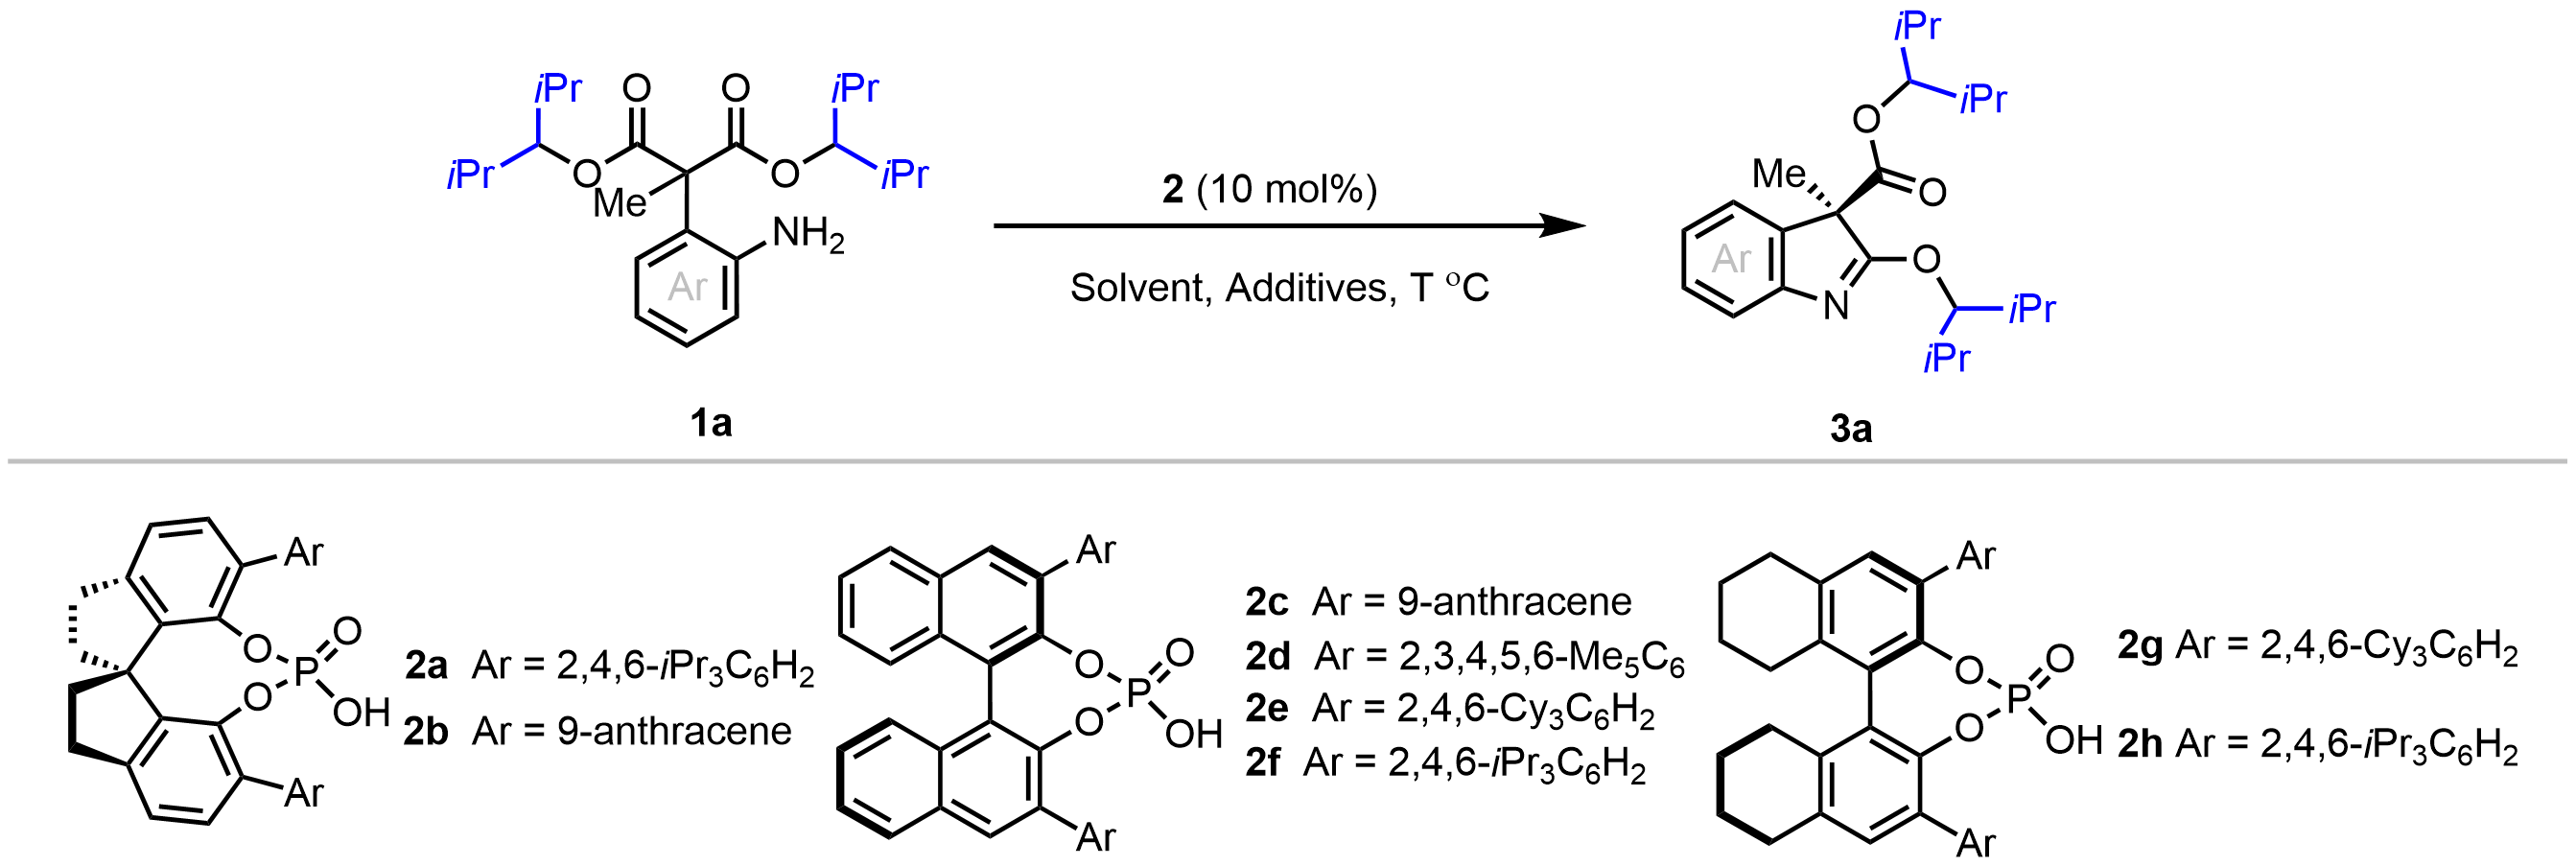
At the beginning, a series of alcohol-derived amino malonic esters were tested for the acid catalyzed ester aminolysis reaction, and we found that the ester moiety derived from 2,4-dimethylpentan-3-ol stands out as the best in terms of enantioselectivity and yield (Table S1). Subsequently, bis(2,4-dimethylpentan-3-yl) 2-(2-aminophenyl)-2-methylmalonate 1a was chosen as the model starting material for the following reaction optimization. The preliminary reactions were performed in CHCl3 (c 0.1 M) at room temperature with chiral phosphoric acids (CPAs)[9,10] as catalysts and 5 Å molecular sieves as additives (Table 1).
Table 1. Screening reaction conditionsa.
| Entry | 2 | Solvent | T (oC) | Additives | 3a (%) | |
| Yieldb | Eec | |||||
| 1 | 2a | CHCl3 | 25 | 5 Å | 8 | 54 |
| 2 | 2b | CHCl3 | 25 | 5 Å | 76 | 83 |
| 3 | 2c | CHCl3 | 25 | 5 Å | 90 | 68 |
| 4 | 2d | CHCl3 | 25 | 5 Å | 91 | 62 |
| 5 | 2e | CHCl3 | 25 | 5 Å | 92 | 85 |
| 6 | 2f | CHCl3 | 25 | 5 Å | 86 | 86 |
| 7 | 2g | CHCl3 | 25 | 5 Å | 91 | 87 |
| 8 | 2h | CHCl3 | 25 | 5 Å | 91 | 89 |
| 9 | 2h | Tol | 25 | 5 Å | 81 | 80 |
| 10 | 2h | DCM | 25 | 5 Å | 66 | 86 |
| 11 | 2h | 1,4-Dixone | 25 | 5 Å | 76 | 86 |
| 12 | 2h | CHCl3 | 25 | 3 Å | 79 | 86 |
| 13 | 2h | CHCl3 | 25 | 4 Å | 84 | 86 |
| 14 | 2h | CHCl3 | 0 | 5 Å | 24 | 89 |
| 15 | 2h | CHCl3 | 40 | 5 Å | 90 | 82 |
a: Standard conditions: 1a (0.05 mmol), 2 (10 mol%), additives (15 mg), solvent (c 0.1 M), sealed tube, argon, 15 h; b: Isolated yields; c: Determined by HPLC analysis; M.S. = molecular sieves; Tol = toluene.
In the beginning, the chiral spirocyclic phosphoric acids 2a-2b were screened. After stirring for 15 h at ambient temperature, the desired dehydrated product 3a was obtained with merely 8% yield with 54% ee, while most of the starting material remained unconsumed. To our delight, catalyst 2b delivered the desired product 3a in 76% yield and 83% ee (entries 1 and 2). In both cases, the alcohol-elimination product obtained in previous literature[6] was also detectable, yet formed in low yields (Table S2). We next investigated a series of binaphthyl-derived phosphoric acids 2c-2f, among which catalysts bearing bulky 2,4,6-triisopropylphenyl and 2,4,6-tricyclohexylphenyl groups both exhibited promising reactivity and enantioselectivity (entries 3-6). Further screening of 8-H-substituted chiral phosphoric acids identified catalyst 2h as the optimal catalyst, affording product 3a in 91% isolated yield and 89% ee (entry 8). With the optimal catalyst identified, we subsequently screened the solvent effect. Chloroform was confirmed as the optimal solvent, superior to toluene, dichloromethane, and 1,4-dioxane, all of which led to diminished yields (entry 8 vs entries 9-11). The impact of molecular sieves was also explored. 5 Å molecular sieves were essential for high yields and enantiocontrol, while 3 Å and 4 Å molecular sieves substantially reduced reaction conversion (entry 8 vs entries 12-13). Temperature optimization revealed that cooling the reaction to 0 °C maintained enantioselectivity but significantly retarded the reaction rate, whereas heating the reaction mixture to 40 °C notably eroded stereochemical control (entries 14-15). Finally, the optimal reaction conditions were identified as follows: 1a (1.0 equiv.), 2h (10 mol%), and 5 Å molecular sieves in chloroform at 25 °C under an argon atmosphere.
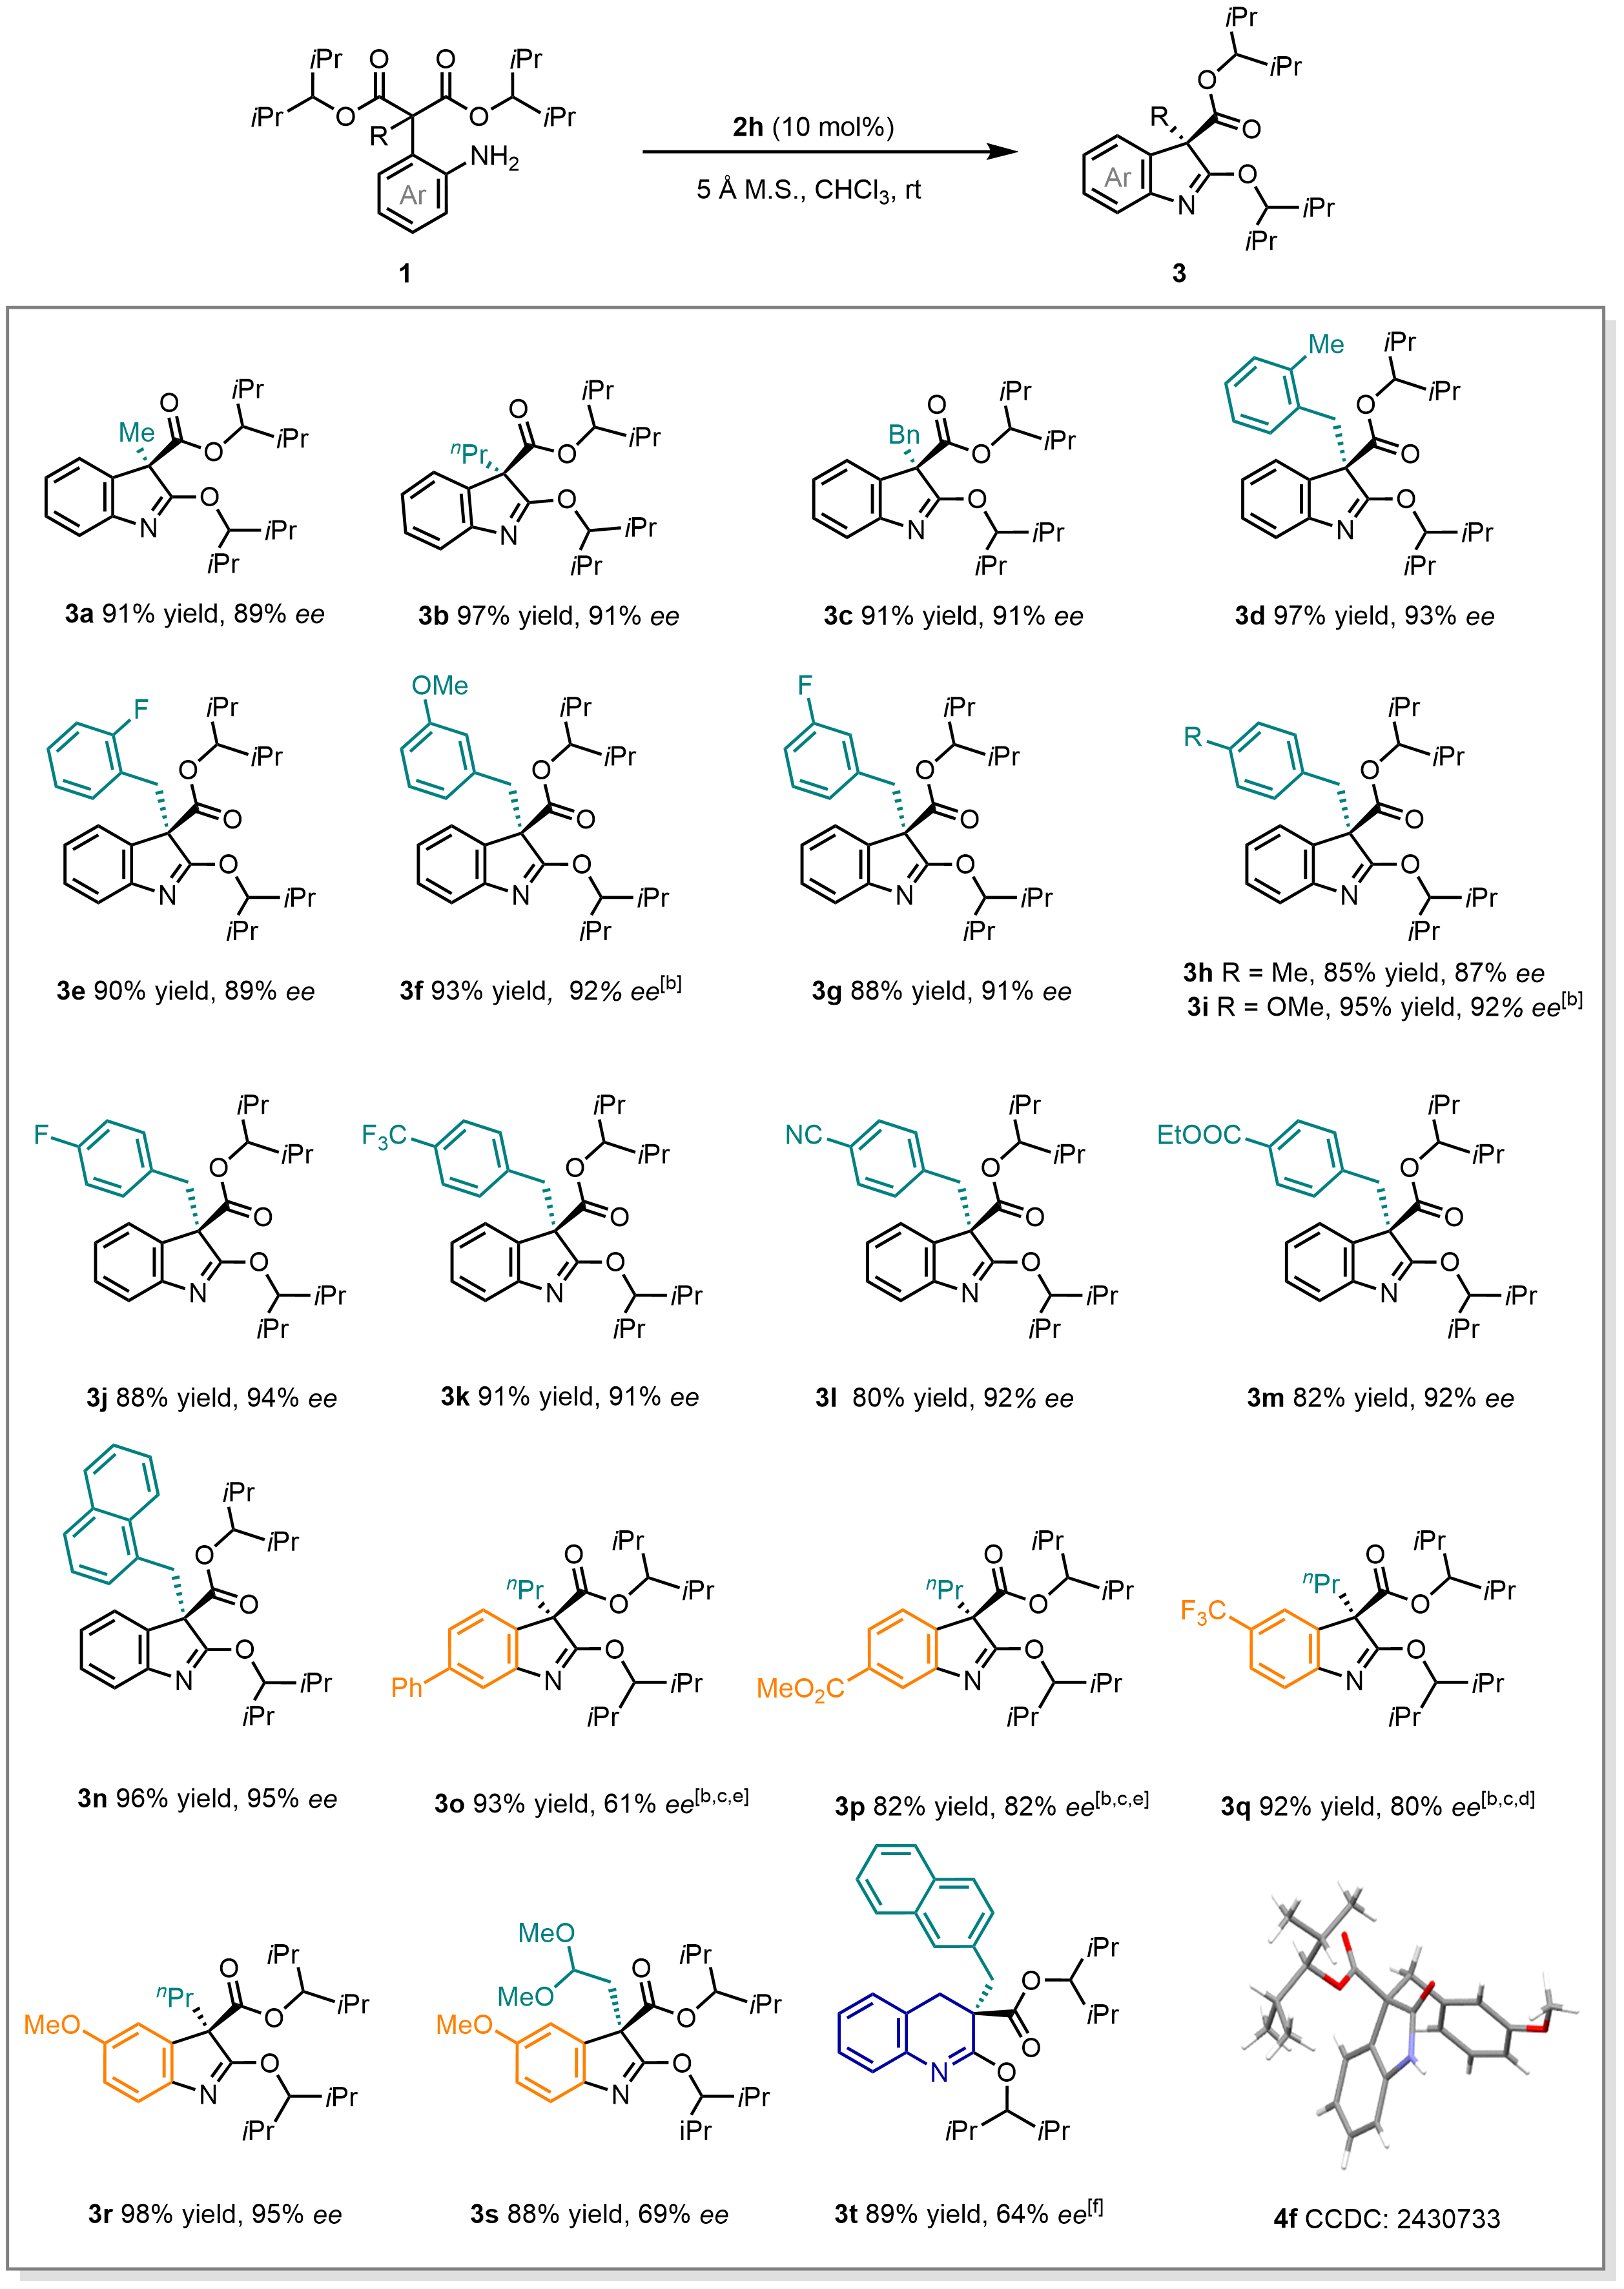
With the optimized conditions in hand, we next explored the substrate scope of this CPA-catalyzed enantioselective desymmetric ester aminolysis (Figure 1). A wide range of 2-aminophenyl malonate derivatives bearing diverse substituents on the quaternary carbon center were well tolerated under the standard conditions, providing the corresponding chiral C3-disubstituted indolenine products in generally high yields and excellent enantioselectivities (3a-3t). Alkyl-substituted substrates, including methyl, n-propyl, and benzyl groups, reacted smoothly to afford products 3a-3c in 91-97% yields with 89-91% ee. Aryl substituents with varying steric and electronic properties were also investigated. Both ortho-substituted (methyl, fluoro) and para-substituted (fluoro, methoxy, cyano, ester, trifluoromethyl) arenes were compatible, delivering the desired products 3d-3m in good to excellent yields (80-97%) and enantioselectivities (87-94% ee). The transformation was also applicable to a naphthyl-substituted substrate, affording product 3n in 96% yield and 95% ee. Furthermore, the substrate scope was expanded to indolenine derivatives bearing phenyl, CO₂Me, CF₃, and OMe substituents (3o-3r), demonstrating good functional group compatibility. Although all these substrates reacted smoothly, diminished enantioselectivity was observed for the phenyl-substituted derivative (3o). Notably, owing to their low polarity, direct separation of the enantiomers of products 3i, 3o-3p via chiral HPLC was not feasible. Hydrolysis was therefore carried out prior to ee analysis. The precursor of Horsfiline[11] 3s was also efficiently prepared under optimized conditions, albeit with a slight drop in enantioselectivity. The diminished enantioselectivity may be attributed to steric hindrance arising from the N,N-dimethyl substituent. Unfavorable steric repulsion between the substrate and catalyst interferes with the formation of critical hydrogen bonds, which are essential for achieving efficient asymmetric catalysis. Furthermore, the corresponding six-membered-ring dihydroquinoline product 3t was smoothly obtained in 89% yield with 64% ee upon prolonging the reaction time to six days under standard conditions.
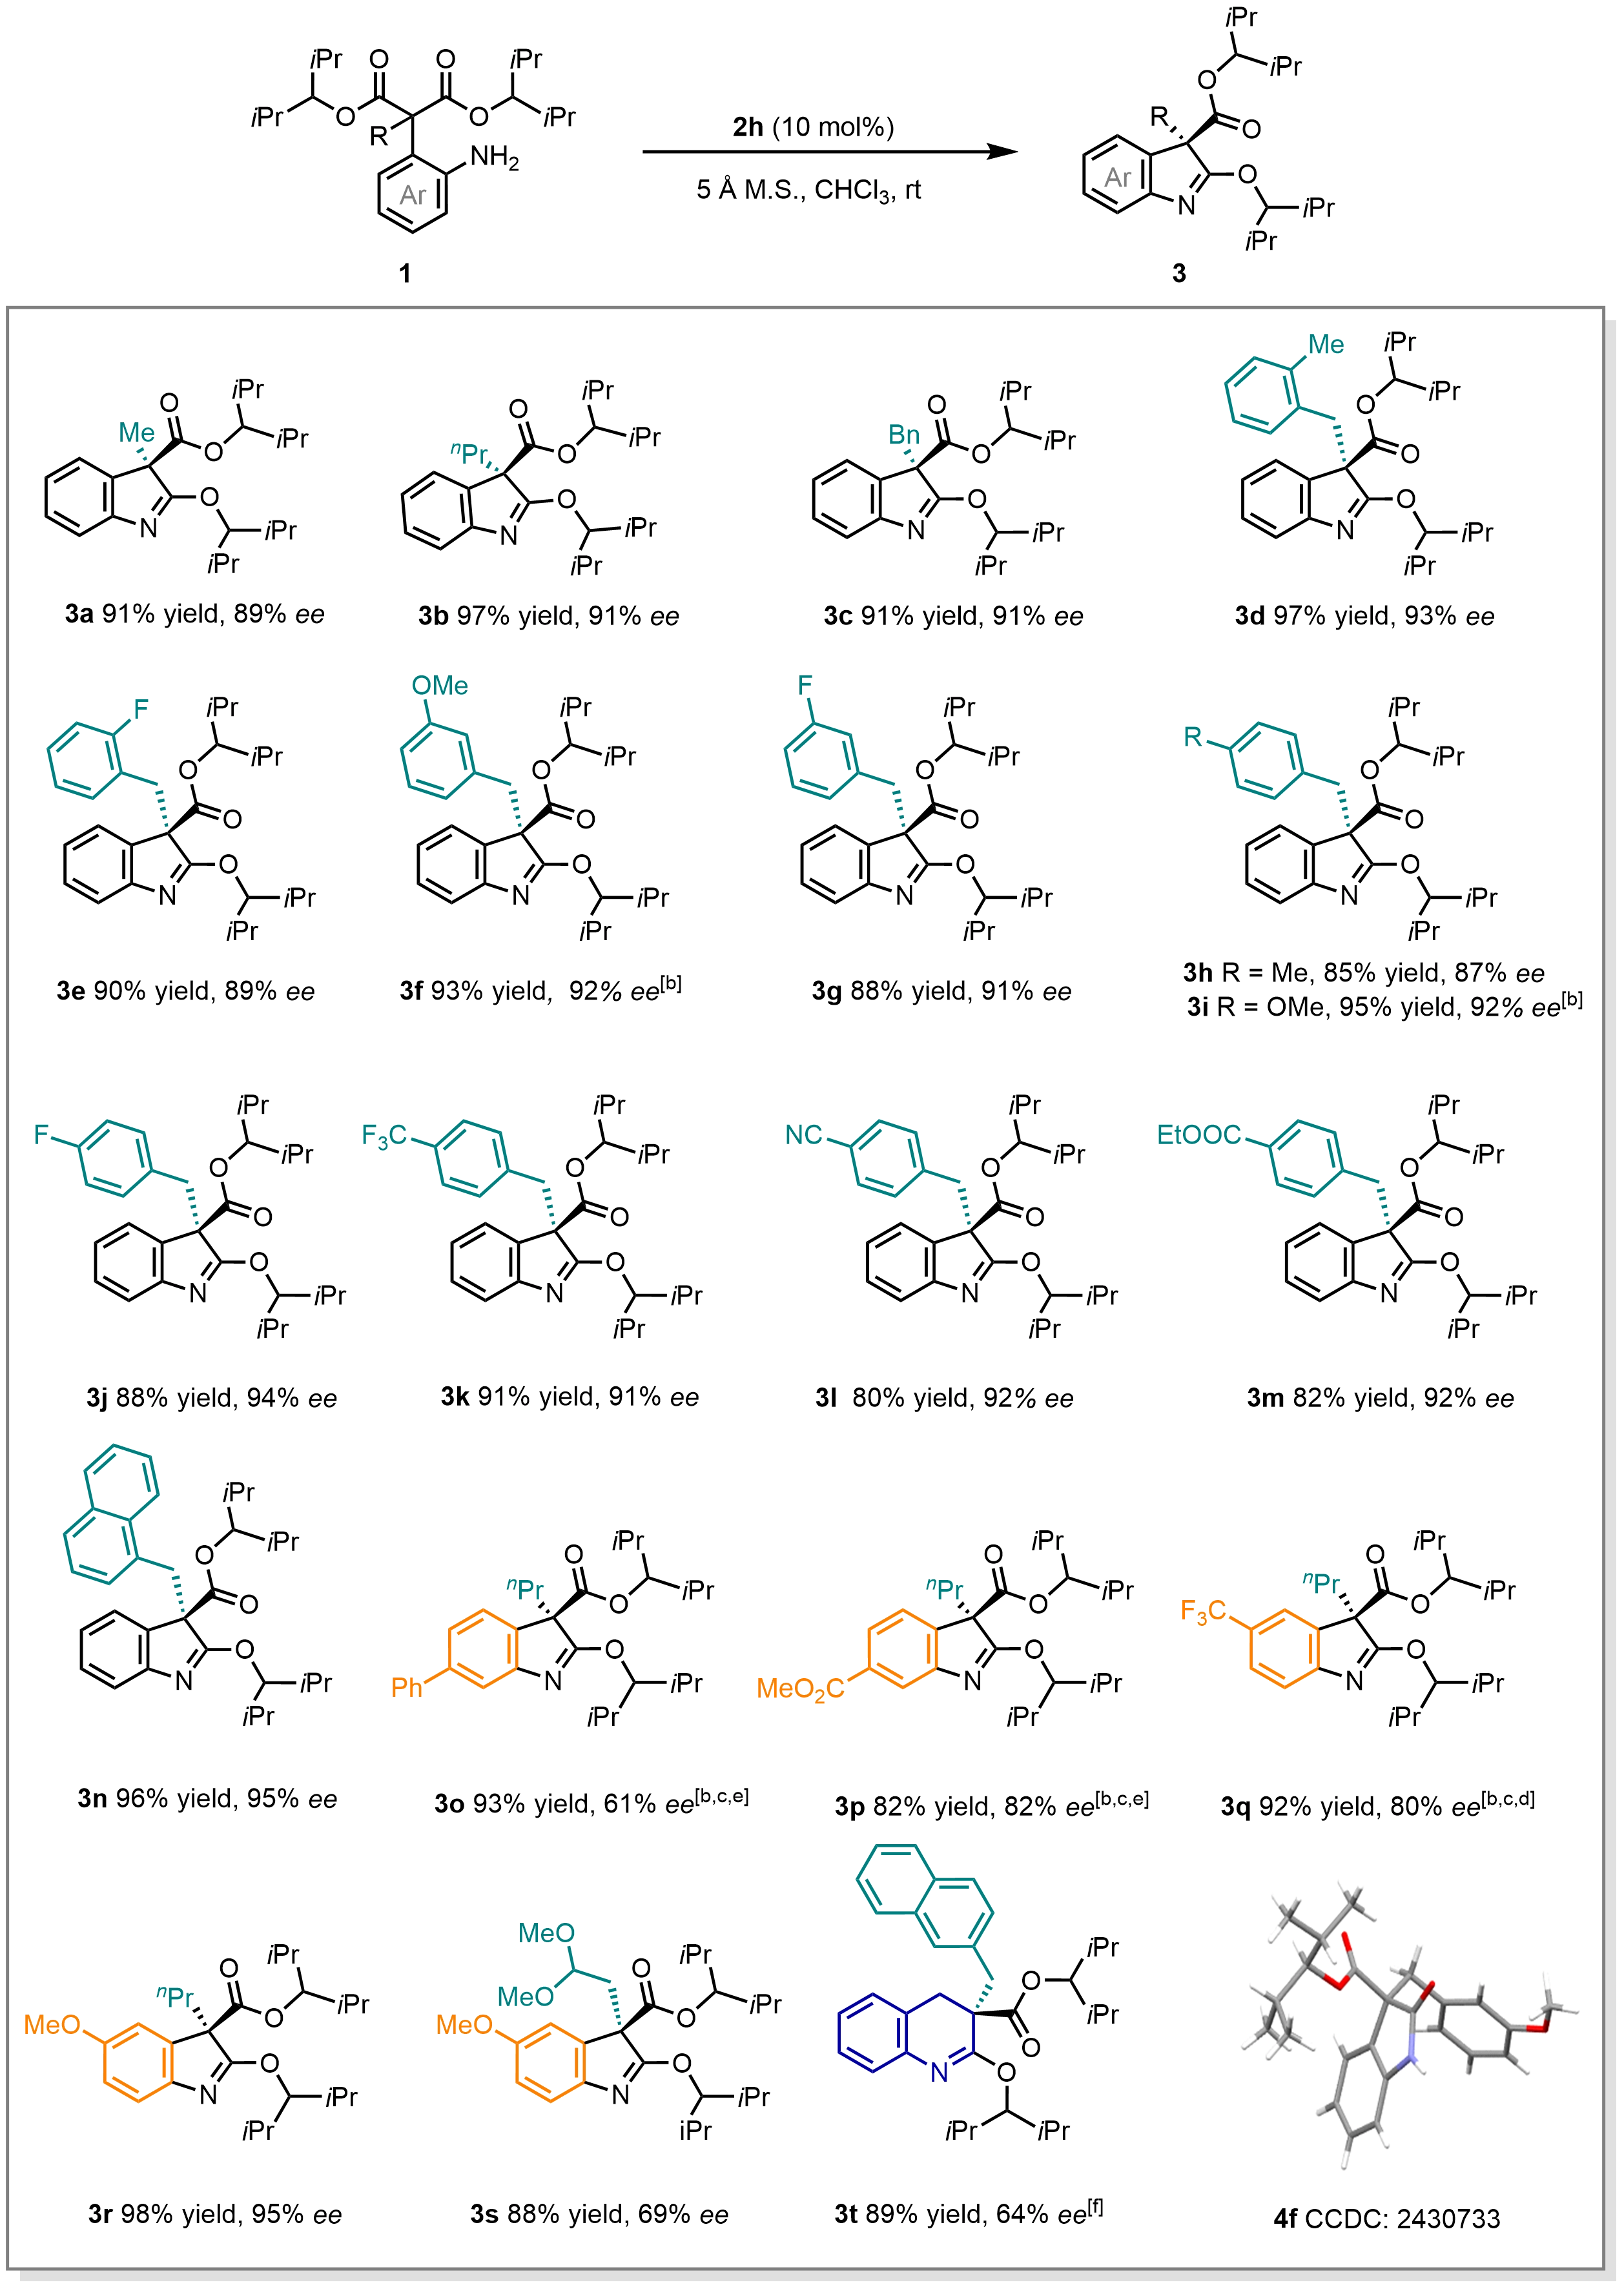
Figure 1. Substrate scope of enantioselective desymmetric ester aminolysisa. a: Conditions: 1 (0.05 mmol), 2h (10 mol%), 5 Å M.S. (15 mg), CHCl3 (c 0.1 M), rt, 15 h, sealed tube, argon; b: Ee value was determined after hydrolysis; c: The reaction temperature was 40 °C; d: 24 h; e:48 h; f: 6 d.
The absolute configuration of 4f (CCDC 2430733) derived from hydrolysis of 3f was determined to be (R) by X-ray crystallography, and the configurations of the other indolenines were assigned accordingly.
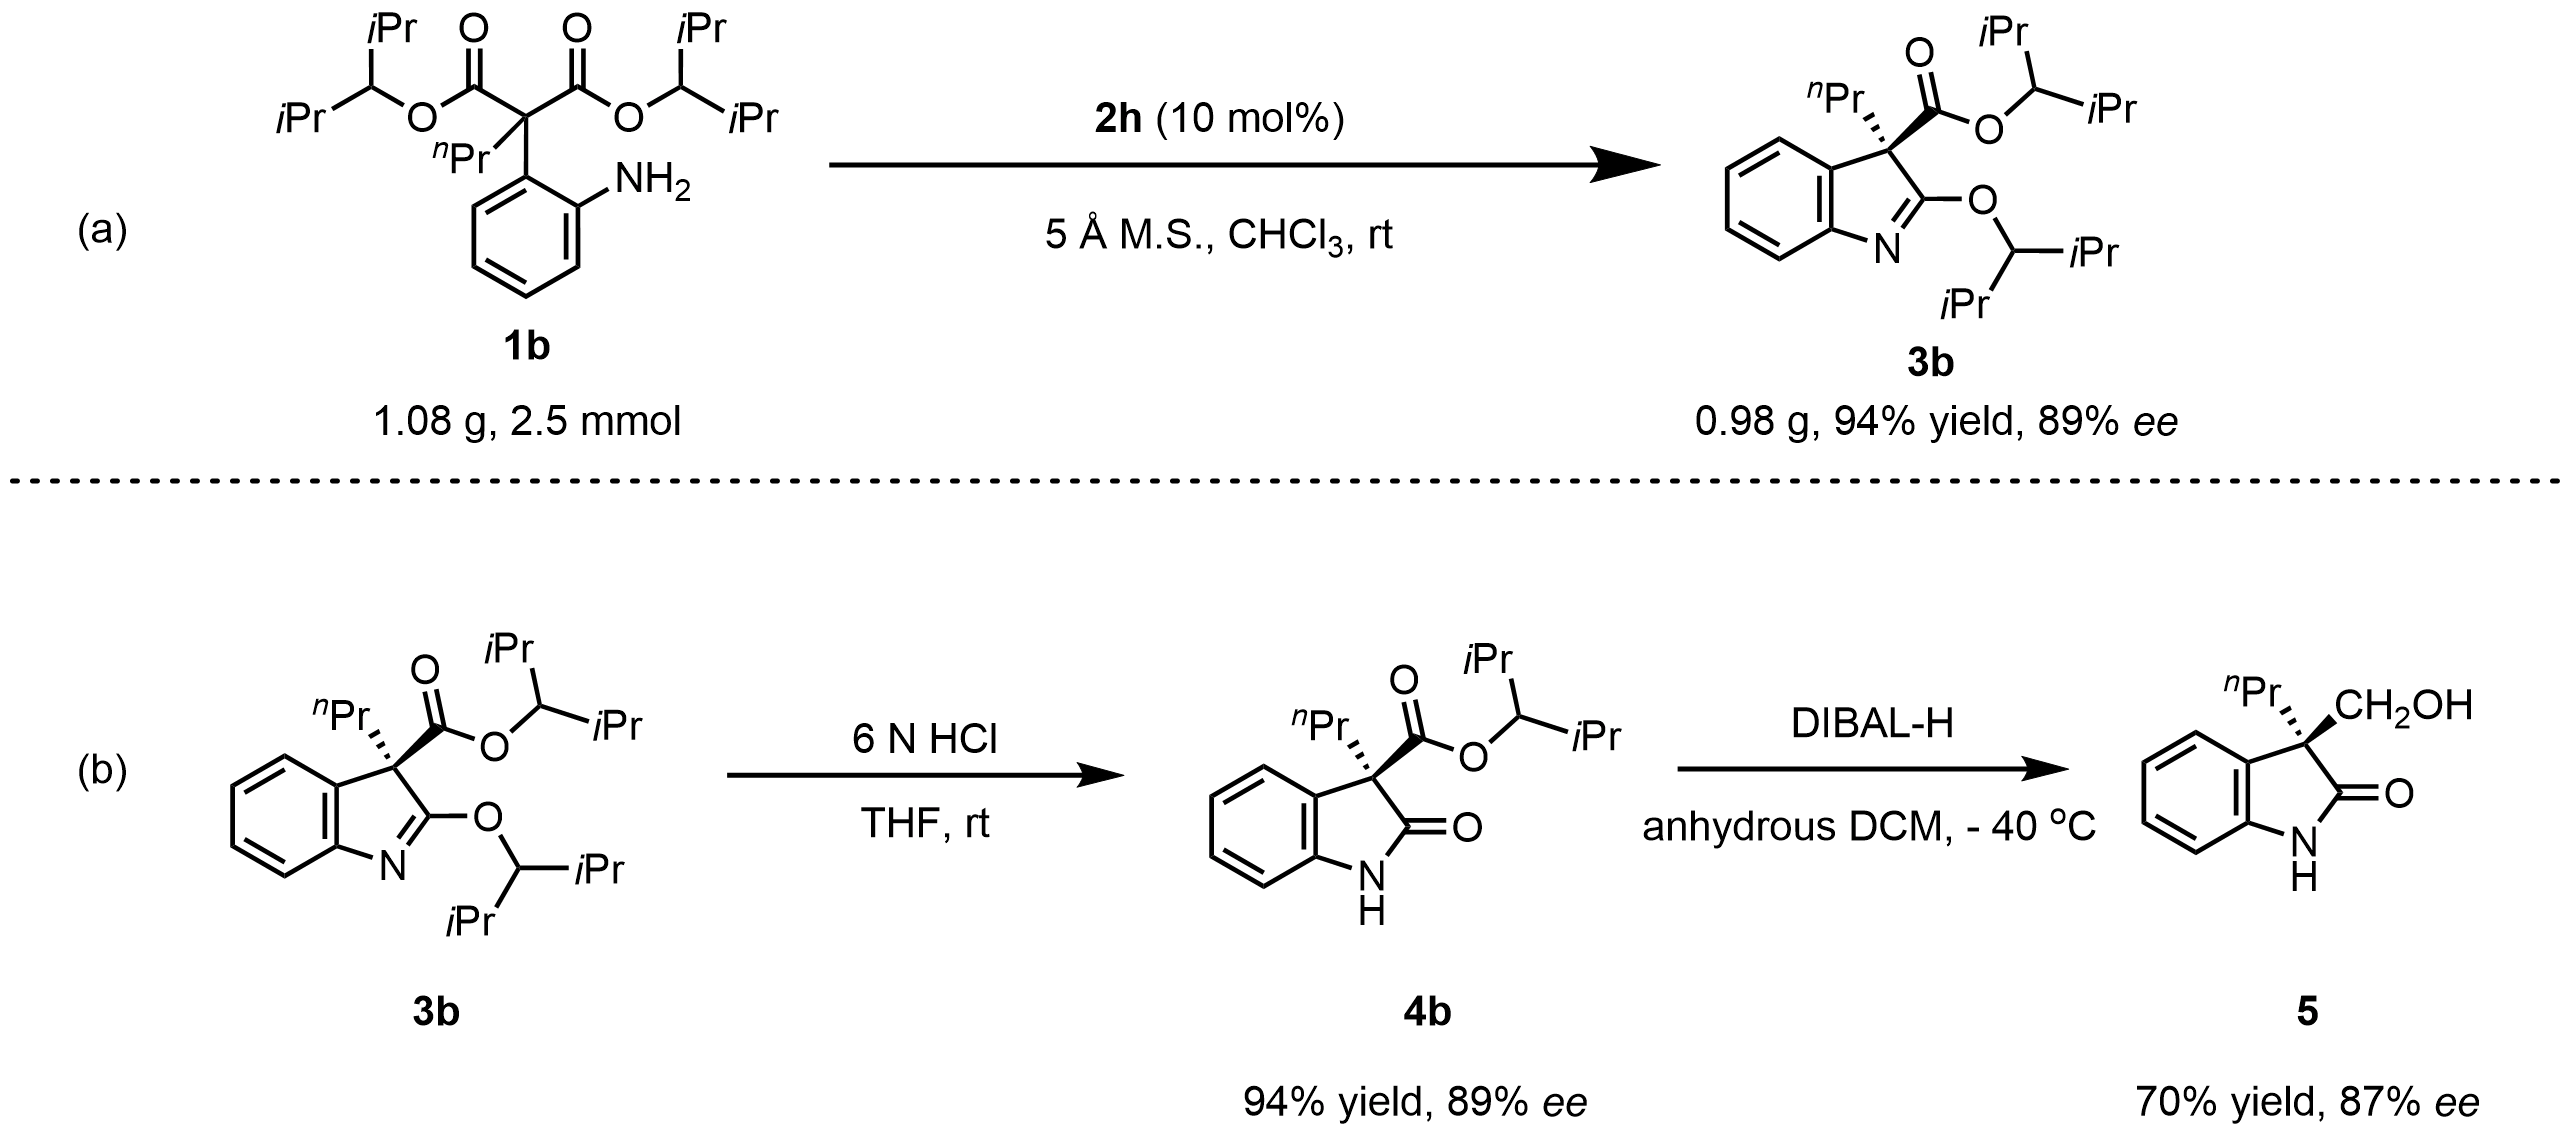
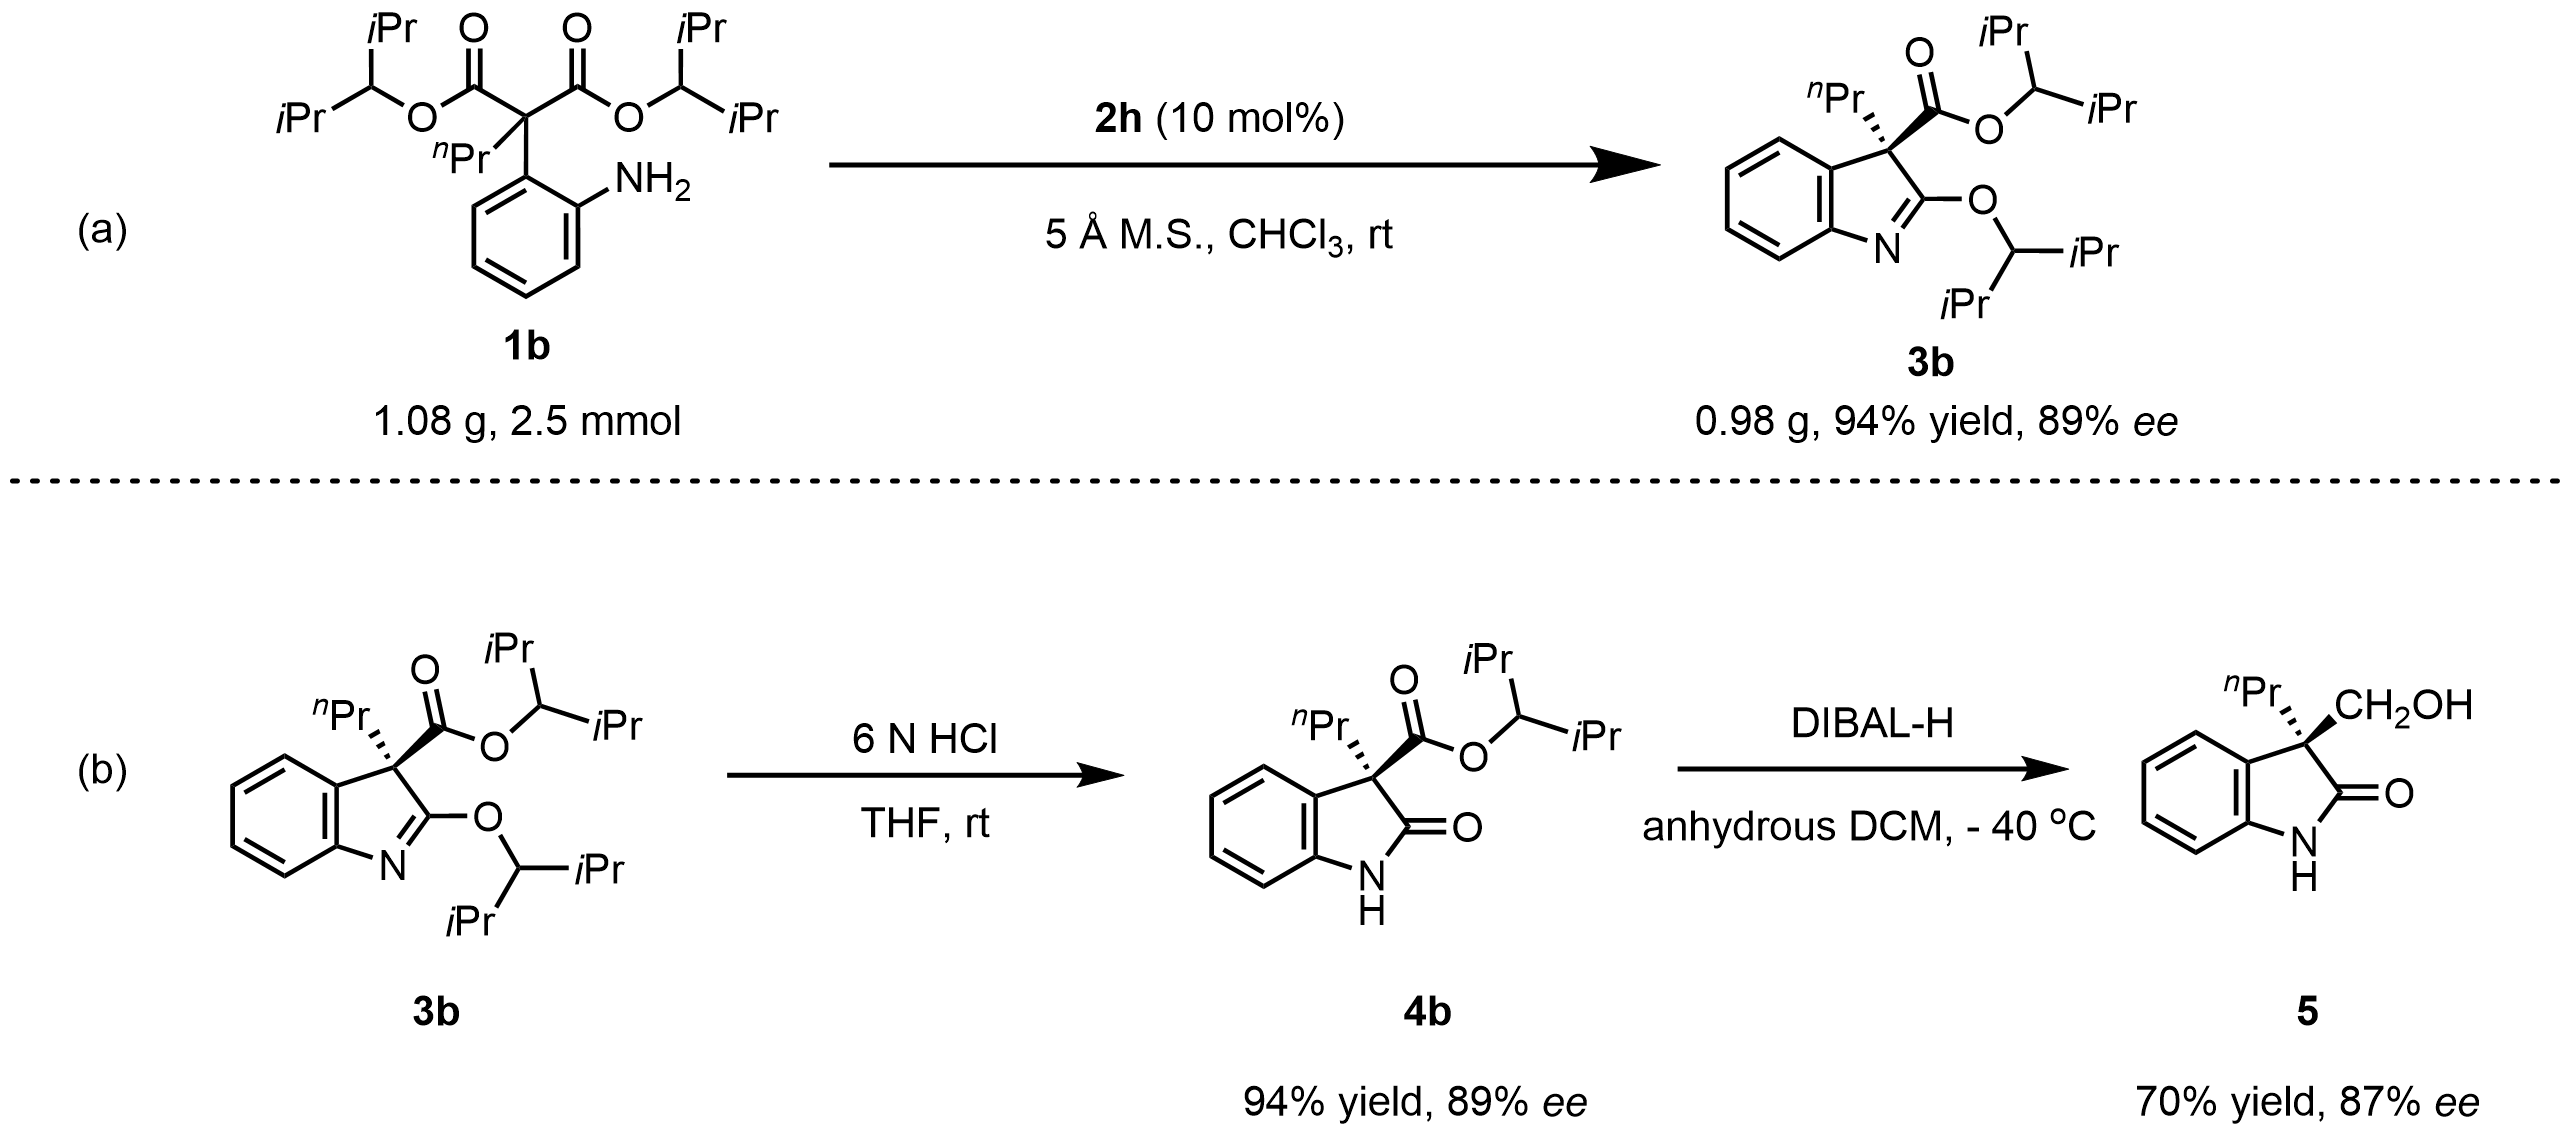
To verify the preparative utility of this catalytic desymmetrization, a gram-scale synthesis of 3b was successfully performed (Figure 2a). The reaction of 2.5 mmol of 1b under the standard conditions proceeded smoothly to deliver product 3b in 94% isolated yield (0.98 g) with 89% ee, demonstrating the good scalability and practicability of this method. Subsequently, we explored further synthetic transformations of the enantioenriched indolenine product 3b (Figure 2b). Treatment of 3b with 6 N HCl in THF at room temperature effected hydrolysis of the C2-isopropoxy group, furnishing the corresponding oxindole derivative 4b in 94% yield with 89% ee. Further reduction of 4b with DIBAL-H in anhydrous DCM at -40 °C afforded the primary alcohol product 5 in 70% yield, albeit with a slight erosion of enantioselectivity (87% ee).
4. Conclusion
In conclusion, we report an organocatalytic enantioselective desymmetrizing intramolecular ester aminolysis to construct chiral C3-disubstituted oxindoles. The reaction follows an unusual dehydration pathway distinct from conventional textbook mechanisms. It tolerates diverse alkyl and aryl substituents and different functional groups. Gram-scale preparation and derivatization verify its practical utility. This method not only offers practical access to enantioenriched oxindoles, but also advances the fundamental understanding of ester aminolysis.
Supplementary materials
The supplementary material for this article is available at: Supplementary materials.
Authors contribution
Zheng WY: Methodology, investigation, data curation, formal analysis, writing-review & editing.
He M: Methodology, investigation, data curation, formal analysis, writing-review & editing.
He YP: Writing-review & editing, supervision, conceptualization, writing-original draft.
Wu H: Supervision, conceptualization, writing-original draft.
Conflicts of interest
The authors declare no conflicts of interest.
Ethical approval
Not applicable.
Consent to participate
Not applicable.
Consent for publication
Not applicable.
Availability of data and materials
The data that support the findings of this study are available from the corresponding author upon reasonable request.
Funding
Financial support for this work was provided by the National Natural Science Foundation of China (Grant No. 22501166).
Copyright
© The Author(s) 2026.
References
-
1. (a) Montalbetti CAGN, Falque V. Amide bond formation and peptide coupling. Tetrahedron. 2005;61(46):10827-10852.[DOI](b) González-Sabín J, Lavandera I, Rebolledo F, Gotor V. Redesigning the mechanism of the lipase-catalysed aminolysis of esters. Tetrahedron Asymmetry. 2006;17(8):1264-1274.[DOI](c) Allen SE, Hsieh SY, Gutierrez O, Bode JW, Kozlowski MC. Concerted amidation of activated esters: Reaction path and origins of selectivity in the kinetic resolution of cyclic amines via N-heterocyclic carbenes and hydroxamic acid cocatalyzed acyl transfer. J Am Chem Soc. 2014;136(33):11783-11791.
-
3. (a) Khetmalis YM, Shivani M, Murugesan S, Chandra Sekhar KVG. Oxindole and its derivatives: A review on recent progress in biological activities. Biomed Pharmacother. 2021;141:111842.
-
4. (a) Zeng XP, Cao ZY, Wang YH, Zhou F, Zhou J. Catalytic enantioselective desymmetrization reactions to all-carbon quaternary stereocenters. Chem Rev. 2016;116(12):7330-7396.(b) Huang Y, Liao JY. Catalytic asymmetric desymmetrization of diesters: Advances and perspectives. Tetrahedron Chem. 2025;14:100134.[DOI]
-
5. (a) Xu P, Huang Z. Catalytic reductive desymmetrization of malonic esters. Nat Chem. 2021;13(7):634-642.[DOI](b) Zheng Y, Zhang S, Low KH, Zi W, Huang Z. A unified and desymmetric approach to chiral tertiary alkyl halides. J Am Chem Soc. 2022;144(4):1951-1961.(c) Xu P, Liu S, Huang Z. Desymmetric partial reduction of malonic esters. J Am Chem Soc. 2022;144(15):6918-6927.(d) Xu P, Shen C, Xu A, Low KH, Huang Z. Desymmetric cyanosilylation of acyclic 1,3-diketones. Angew Chem Int Ed. 2022;61(37):e202208443.(e) Liu H, Lau VHM, Xu P, Chan TH, Huang Z. Diverse synthesis of α-tertiary amines and tertiary alcohols via desymmetric reduction of malonic esters. Nat Commun. 2022;13(1):4759.(f) Zheng WF, Chen J, Qi X, Huang Z. Modular and diverse synthesis of amino acids via asymmetric decarboxylative protonation of aminomalonic acids. Nat Chem. 2023;15(12):1672-1682.[DOI](g) Qabaja G, Wilent JE, Benavides AR, Bullard GE, Petersen KS. Facile synthesis of versatile enantioenriched α-substituted hydroxy esters through a Brønsted acid catalyzed kinetic resolution. Org Lett. 2013;15(6):1266-1269.[DOI](h) Wilent J, Petersen KS. Enantioselective desymmetrization of diesters. J Org Chem. 2014;79(5):2303-2307.[DOI](i) Puglisi A, Giustini C, Ricucci A, Perotti E, Massaro L, Morra D, et al. Synthesis of benzofuranones via malonates desymmetrization: Yield increase by the portion-wise addition of quinones. Chemistry. 2018;24(27):6941-6945.(j) Kelley AM, Minerali E, Wilent JE, Chambers NJ, Stingley KJ, Wilson GT, et al. Asymmetric synthesis of novel spirocycles via a chiral phosphoric acid catalyzed desymmetrization. Tetrahedron Lett. 2019;60(18):1262-1264.[DOI](k) Kelley AM, Haywood RD, White JC, Petersen KS. Enantioselective desymmetrizations of diesters to synthesize fully substituted chiral centers of 3,4-dihydrocoumarins and related compounds. ChemistrySelect. 2020;5(10):3018-3022.[DOI](l) Frost JA, Korb SM, Green FE, Youngblood KC, Petersen KS. Chiral Brønsted acid catalyzed cascade alcohol deprotection and enantioselective cyclization. ACS Omega. 2024;9(2):2962-2969.[DOI]
-
6. Xie C, Kim J, Mai BK, Cao S, Ye R, Wang XY, et al. Enantioselective synthesis of quaternary oxindoles: Desymmetrizing Staudinger–Aza-Wittig reaction enabled by a bespoke HypPhos oxide catalyst. J Am Chem Soc. 2022;144(46):21318-21327.[DOI]
-
7. Sumiyoshi T, Ishida K, Shimizu M, Yamai YS, Natsutani I, Uesato S, et al. Asymmetric synthesis of t-butyl 3-alkyl-oxindole-3-carboxylates via chiral phosphoric acid catalyzed desymmetrization of di-t-butyl 2-alkyl-2-(2-aminophenyl)malonates. Heterocycles. 2019;99(2):1398.[DOI]
-
8. (a) He YP, Li ZC, Wang ZQ, Zheng WY, Wu H. Enamine acylation enabled desymmetrization of malonic esters. J Am Chem Soc. 2024;146(38):26387-26396.(b) Zheng WY, Wang ZQ, Li XZ, Li ZC, Wu H, Rexit AA, et al. Enantioselective catalytic Urech hydantoin synthesis. Org Chem Front. 2025;12(16):4417-4423.[DOI]
Copyright
© The Author(s) 2027. This is an Open Access article licensed under a Creative Commons Attribution 4.0 International License (https://creativecommons.org/licenses/by/4.0/), which permits unrestricted use, sharing, adaptation, distribution and reproduction in any medium or format, for any purpose, even commercially, as long as you give appropriate credit to the original author(s) and the source, provide a link to the Creative Commons license, and indicate if changes were made.
Publisher’s Note
Science Exploration remains a neutral stance on jurisdictional claims in published
maps
and institutional affiliations. The views expressed in this article are solely those
of
the author(s) and do not reflect the opinions of the Editors or the publisher.
Share And Cite


Science Exploration Style
Zheng WY, He M, Wu H, He YP. Enantioselective catalytic desymmetric ester aminolysis. Chiral Chem. 2027;3:202628. https://doi.org/10.70401/cc.2026.0037
Tips
Copy completed.
Submit a Manuscript
Author Instructions
Cite this Article
Article Metrics
0
View
0
Download
Cited
Article Updates

 Get citation
Science Exploration Style

Zheng WY, He M, Wu H, He YP. Enantioselective catalytic desymmetric ester aminolysis. Chiral Chem. 2027;3:202628. https://doi.org/10.70401/cc.2026.0037
copy
Share Link
copy